
Both NDA and ANDA are the regulatory pathways for pharmaceutical companies to obtain FDA approval to market drugs in the US
In NDA and ANDA, NDA stands for ‘New Drug Application’ and ANDA stands for ‘Abbreviated New Drug Application’.
Both NDA and ANDA are the regulatory pathways for pharmaceutical companies to obtain FDA approval to market drugs in the US. NDA is used for new, previously unapproved drugs, and the ANDA is for generic versions of drugs already on the market
An NDA is a formal proposal submitted to the U.S. Food and Drug Administration (FDA) requesting approval to market a new drug in the United States. It is the regulatory pathway for innovator drugs—medicines that contain new active ingredients or new combinations of existing drugs.
Key Points:
An ANDA is the application used to seek approval to market a generic version of an already approved drug (the reference listed drug or RLD). It is called “abbreviated” because it does not require the generic drug sponsor to submit preclinical and clinical data on safety and effectiveness again.
Key Points:
| Aspect | NDA | ANDA |
|---|---|---|
| Purpose | Secures approval for a novel, unapproved drug | Approval for generic drugs |
| Data Required | Full safety & efficacy data | Bioequivalence data only |
| Clinical Data Requirements | Requires extensive preclinical (animal) testing and three comprehensive phases of human clinical trials to prove safety and efficacy | Bypasses original animal and human clinical trials, requiring instead scientific proof of bioequivalence (matching the rate and extent of absorption of the active |
| Patent Consideration | Initial patent and exclusivity | Must address existing patents of RLD |
| Review Process | More extensive & lengthy | Usually shorter & less expensive |
| Chemistry, Manufacturing, and Controls (CMC) | Must provide full details on manufacturing, synthesis, processing, and packaging. | Must match the reference drug’s active ingredients, dosage form, strength, route of administration, and quality standard |
| Timeline and Cost | Highly expensive and lengthy, often taking 10 to 15 years from development to final approval. | Faster and less expensive, with standard reviews typically taking about 8 to 10 months |
| Market Exclusivity | Usually grants 5 years of market exclusivity for new chemical entities. | May grant 180 days of generic market exclusivity for the first applicant to successfully file a Paragraph IV certification |
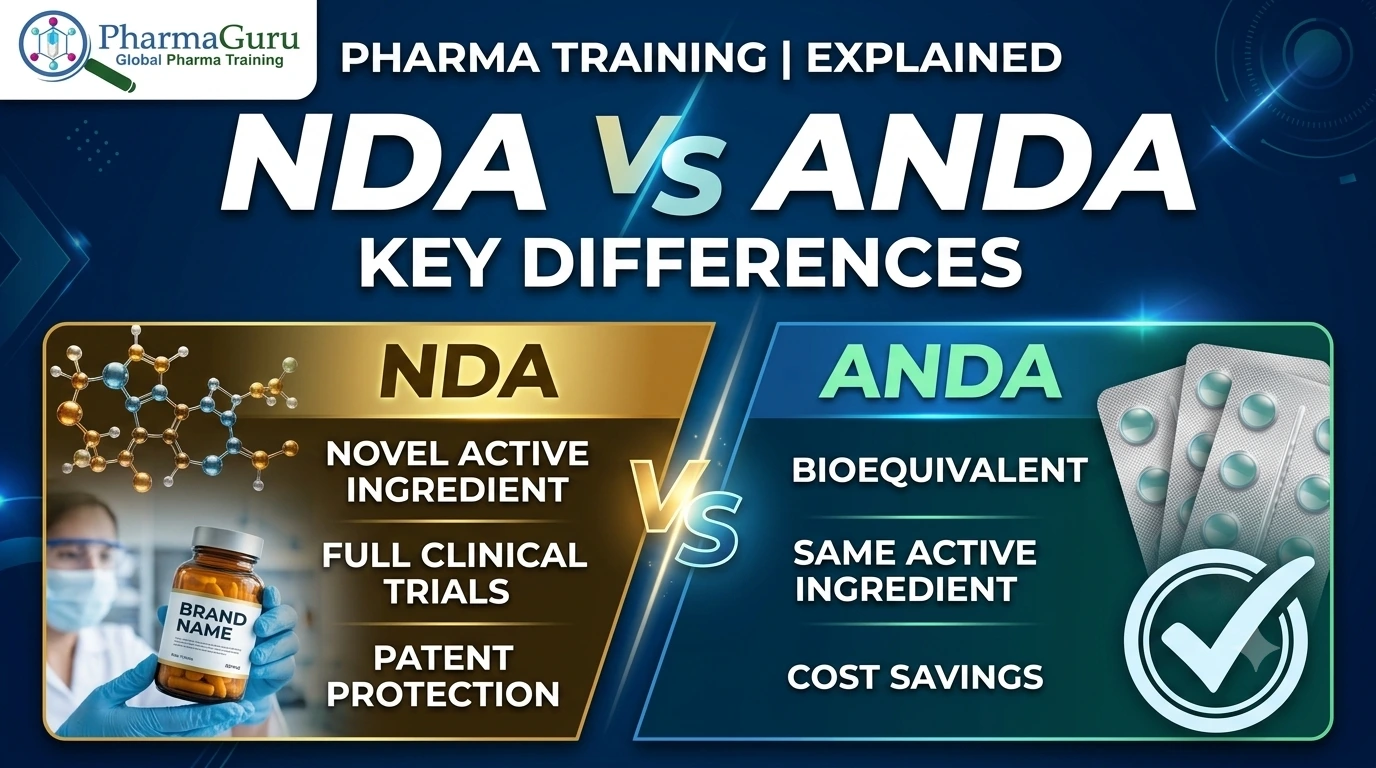
Understanding the distinction between NDA and ANDA is fundamental for anyone working in the pharmaceutical or healthcare industry. While the NDA process is designed to bring novel therapies to patients, the ANDA process allows for cost-effective generic alternatives, helping make medicines accessible to a wider population.
Both play critical roles in the lifecycle of drug development and approval, balancing innovation with affordability.
You May Like
The NDA and ANDA processes are both regulatory pathways for pharmaceutical companies to obtain FDA approval to market drugs in the US, with the NDA used for new, previously unapproved drugs, and the ANDA for generic versions of drugs already on the market.
NDA is used for new, previously unapproved drugs, and the ANDA is for generic versions of drugs already on the market.
An NDA (New Drug Application) is a submission to the FDA requesting approval to market a new drug, including comprehensive data on its safety, effectiveness, and manufacturing.
No, ANDA applicants must demonstrate bioequivalence but do not need to conduct full clinical trials like an NDA.
The NDA approval process typically takes 10 months under the standard review, but it can be shorter for priority reviews.
New Drug Application
Abbreviated New Drug Application
Further Reading
Quick Links